
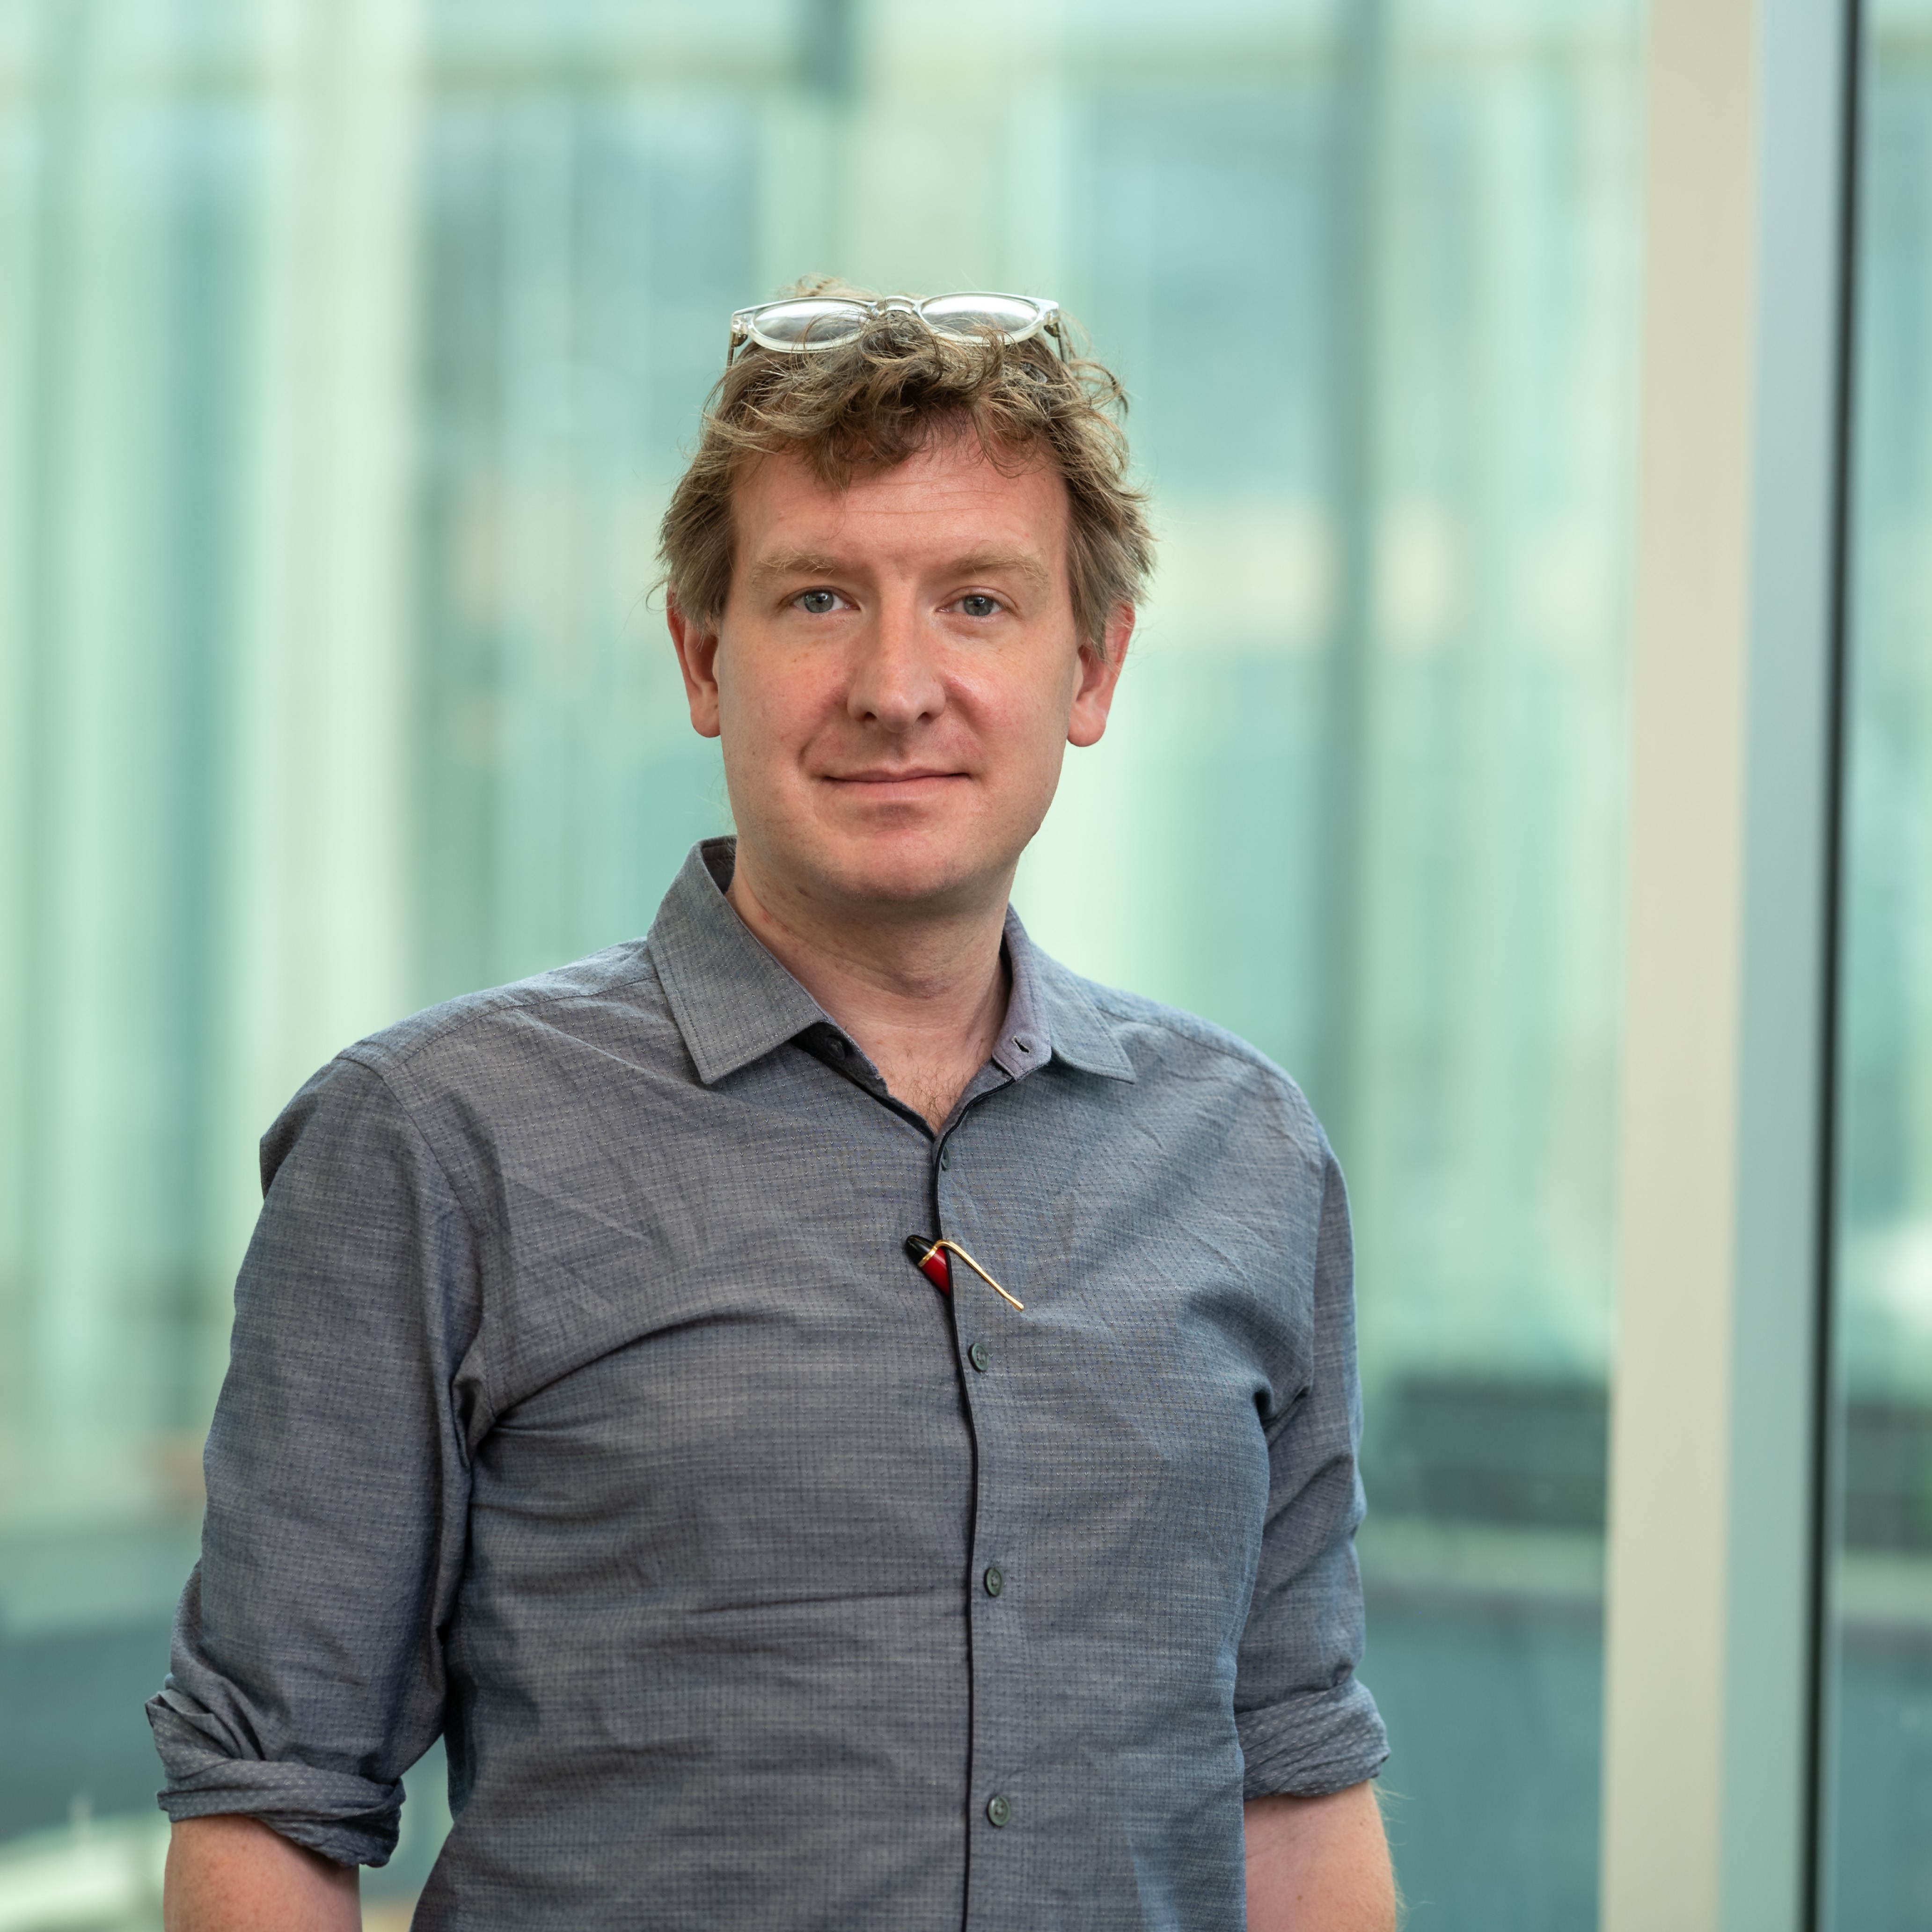
Carl Kingsford
· Herbert A. Simon Professor and Co-Director of the Joint Carnegie Mellon-University of Pittsburgh Ph.D Program in Computational BiologyCarnegie Mellon University · Ray and Stephanie Lane Computational Biology Department
Active 2000–2026
Academic metrics are sourced from OpenAlex and public funding records; values may differ from Google Scholar.
About
Carl Kingsford is the Herbert A. Simon Professor of Computer Science in the Ray and Stephanie Lane Computational Biology Department at Carnegie Mellon University. He is recognized as a trailblazer in computational molecular biology, showcasing sustained innovation in scalable algorithmic approaches. His research focuses on the development of computational methods and algorithms for biological data analysis, including genome graph construction, sequence analysis, and the study of genomic variation. Kingsford's contributions have significantly advanced the understanding of genome structure and function through algorithmic innovations, and he has been honored as an ISCB Fellow for his impactful work in the field.
Research topics
- Computer Science
- Biology
- Artificial Intelligence
- Computational biology
- Data Mining
- Anatomy
- Cell biology
- Algorithm
Selected publications
Alignment and mapping methodology influence transcript abundance estimation
Genome biology · 2020 · 192 citations
BACKGROUND: The accuracy of transcript quantification using RNA-seq data depends on many factors, such as the choice of alignment or mapping method and the quantification model being adopted. While the choice of quantification model has been shown to be important, considerably less attention has been given to comparing the effect of various read alignment approaches on quantification accuracy. RESULTS: We investigate the influence of mapping and alignment on the accuracy of transcript quantifica…
Advances and prospects for the Human BioMolecular Atlas Program (HuBMAP)
Nature Cell Biology · 2023 · 170 citations
Improved design and analysis of practical minimizers
Bioinformatics · 2020 · 67 citations
Abstract Motivation Minimizers are methods to sample k-mers from a string, with the guarantee that similar set of k-mers will be chosen on similar strings. It is parameterized by the k-mer length k, a window length w and an order on the k-mers. Minimizers are used in a large number of softwares and pipelines to improve computation efficiency and decrease memory usage. Despite the method’s popularity, many theoretical questions regarding its performance remain open. The core metric for measuring…
How Much Data Is Sufficient to Learn High-Performing Algorithms?
Journal of the ACM · 2024 · 13 citations
Algorithms often have tunable parameters that impact performance metrics such as runtime and solution quality. For many algorithms used in practice, no parameter settings admit meaningful worst-case bounds, so the parameters are made available for the user to tune. Alternatively, parameters may be tuned implicitly within the proof of a worst-case approximation ratio or runtime bound. Worst-case instances, however, may be rare or nonexistent in practice. A growing body of research has demonstrate…
<i>k</i> -nonical space: sketching with reverse complements
Bioinformatics · 2024-10-17 · 7 citations
articleOpen accessSenior authorMOTIVATION: Sequences equivalent to their reverse complements (i.e. double-stranded DNA) have no analogue in text analysis and non-biological string algorithms. Despite this striking difference, algorithms designed for computational biology (e.g. sketching algorithms) are designed and tested in the same way as classical string algorithms. Then, as a post-processing step, these algorithms are adapted to work with genomic sequences by folding a k-mer and its reverse complement into a single sequen…
Recent grants
Algorithms for Managing Uncertainty in Chromosome Conformation Capture Data
NIH · $1.3M · 2013–2017
CAREER: Model-based Reconstruction of Ancient Biological Networks
NSF · $177k · 2011–2012
NIH · $1.2M · 2017–2022
Frequent coauthors
- 42 shared
Rob Patro
University of Maryland, College Park
- 38 shared
Guillaume Marçais
Carnegie Mellon University
- 25 shared
Mingfu Shao
Pennsylvania State University
- 21 shared
Dan DeBlasio
Carnegie Mellon University
- 19 shared
Cong Ma
Northwestern Polytechnical University
- 19 shared
Geet Duggal
DNAnexus (United States)
- 17 shared
Charlotte Soneson
SIB Swiss Institute of Bioinformatics
- 16 shared
Darya Filippova
Labs
Similar researchers at Carnegie Mellon University
- Resume-aware match score
- Save to shortlist
- AI-drafted outreach
See your match with Carl Kingsford
PhdFit ranks faculty by your research interests, methods, and publications — grounded in their actual work, not templates.
- Free to start
- No credit card
- 30-second signup